Research
My research develops computational methods for understanding molecules and materials. Across electronic-structure theory, quantum computing, and machine learning, I seek representations that turn physical insight into accurate and efficient simulations.
Stochastic Tensor Contraction
Accurate electronic-structure calculations are essential for predicting reaction energies, understanding chemical bonding, and describing molecules and materials. Much of this accuracy comes from treating electronic correlation—the coordinated motion of electrons beyond a mean-field description. Coupled-cluster methods, particularly CCSD(T), provide a reliable treatment for many chemical systems, but their steep computational cost limits the size of systems that can be studied.
The locality of electronic correlation offers a route to lower cost. Conventional local-correlation methods restrict the correlation contributions included in a calculation, neglecting sufficiently weak or distant ones. These approaches are highly successful, but controlling the neglected contributions becomes more difficult when correlation is spatially extended or exceptionally high accuracy is required. The challenge is to exploit the uneven importance of correlation contributions while retaining their collective effect.
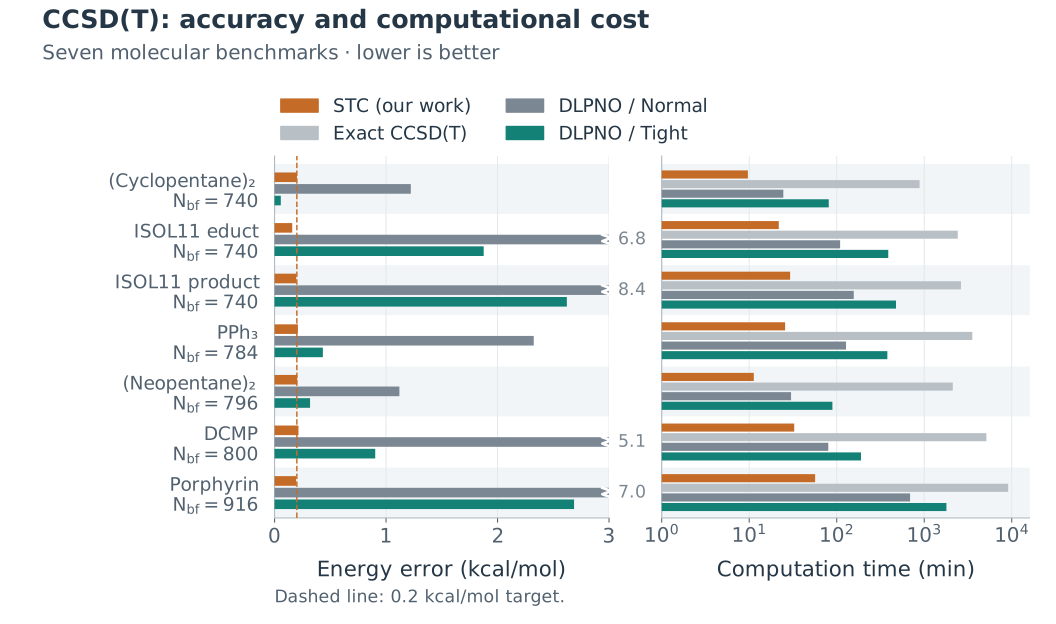
Our stochastic tensor contraction (STC) framework addresses this challenge through statistical sampling. Correlated electronic-structure methods involve expensive tensor contractions: sums over many combinations of orbital indices. STC estimates these sums using samples chosen to reflect the structure of their contributions. Locality concentrates sampling effort on the most important contributions, while distant effects remain included. Computational effort is governed by the cost and statistical variance of this estimation, opening a different route to efficient correlated calculations.
With Garnet Chan, I applied STC to CCSD(T), achieving simultaneous order-of-magnitude improvements in efficiency and accuracy over state-of-the-art local-correlation methods in the reported benchmarks, with reduced sensitivity to electronic delocalization. In subsequent work on MP2 exchange, we combined deterministic evaluation with grouped stochastic sampling to reduce the computational prefactor. The exchange estimator is unbiased, and its sampling cost can be planned for a specified statistical error. Together, these studies establish STC as a tensor-contraction framework with applications across correlated quantum chemistry.
Related publications
- Stochastic tensor contraction for quantum chemistry. PNAS 123, e2609149123 (2026).
- Stochastic tensor contraction for efficient MP2 exchange. Preprint (2026).
Quasiparticle Quantum Simulation
Quantum computers offer a route to electronic-structure problems that are difficult for classical methods, but directly simulating a realistic material’s full electronic Hamiltonian can require enormous fault-tolerant resources. Classical calculations often succeed by using effective descriptions that incorporate much of the electronic background and focus explicitly on the excitations relevant to a particular observable. These descriptions offer a physical starting point for quantum algorithm design.
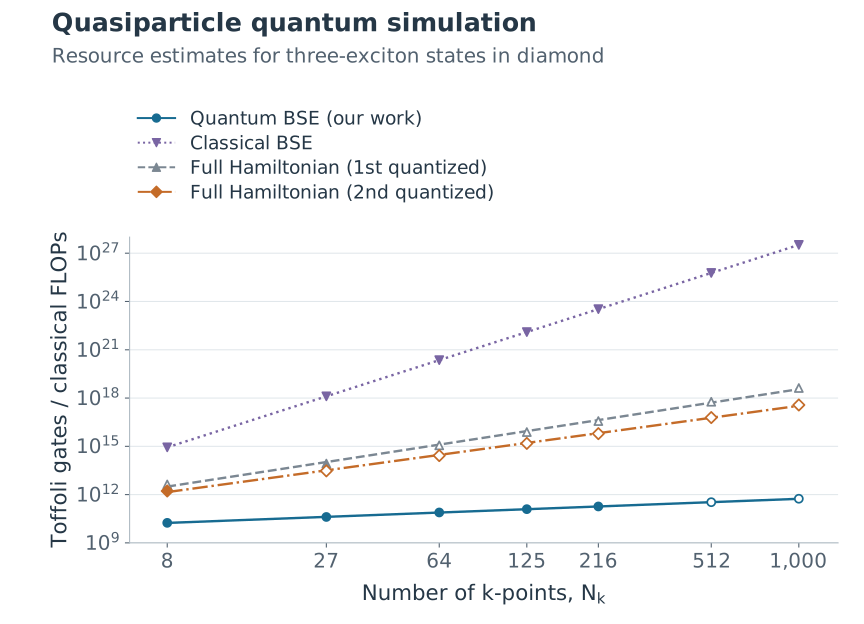
With Jielun Chen and Garnet Chan, I developed quasiparticle quantum simulation of multiexciton states using the Bethe–Salpeter equation (BSE). In this description, excited electrons and the holes they leave behind interact within an effective theory that accounts for the surrounding electronic system. We constructed an orbital-based quantum encoding of this problem and reduced its implementation cost using integral factorization and crystal symmetries. The quantum calculation solves the BSE excitation problem, with accuracy determined in part by the underlying effective-theory approximations.
For representative three-exciton energy calculations in semiconductor models of roughly a thousand atoms, our resource estimates show six to seven orders of magnitude fewer Toffoli gates than full-Hamiltonian simulation estimates. An alternative implementation prioritizing qubit economy requires only a few hundred logical qubits. These results demonstrate how physically motivated classical descriptions can guide quantum algorithms toward realistic materials problems.
Related publication
Quantum-information Inspired Computational Tools
Clifford methods and stabilizer ground states
Stabilizer states and Clifford operations have compact classical descriptions, making them useful tools for designing both quantum and classical algorithms. Our Clifford-based Hamiltonian engineering for molecules (CHEM) uses a Hamiltonian transformation to give shallow variational circuits a Hartree–Fock starting energy and improve their initial energy gradients without adding quantum resources. In related work, we developed stabilizer ground-state algorithms, including an exact, linear-scaling algorithm for one-dimensional local Hamiltonians. Here, “ground state” denotes the lowest-energy state within the stabilizer family. These states provide tractable approximations and starting points for more elaborate many-body methods.
- Towards chemical accuracy with shallow quantum circuits: A Clifford-based Hamiltonian engineering approach. J. Chem. Theory Comput. 20, 695–707 (2024).
- Stabilizer ground states for simulating quantum many-body physics: theory, algorithms, and applications. Quantum 9, 1782 (2025).
Shadow tomography for classical tensor networks
A compact representation of a many-body state does not necessarily make its observables inexpensive to compute. Long-range Hamiltonians, in particular, involve many terms whose evaluation can dominate a tensor-network calculation. Inspired by shadow tomography in quantum measurement, our work adapts randomized estimators to the contraction requirements of classical tensor networks. We developed strategies for both spin and fermionic systems that improve sampling efficiency and yield more stable observable gradients in variational optimization. Demonstrations include a long-range Heisenberg model and an ab initio quantum-chemistry Hamiltonian, showing how quantum-measurement ideas can improve entirely classical calculations.
- Shadow tomography for classical tensor network simulations. Preprint (2026).
Molecular-Orbital-Based Machine Learning
High-accuracy molecular calculations are often too expensive to repeat across the many structures needed for reaction pathways, molecular dynamics, or chemical screening. Machine learning can reduce this cost by learning from a smaller set of accurate reference calculations. A central question is how to represent a molecule so that the model can transfer what it learns across changes in size, geometry, and chemical environment.
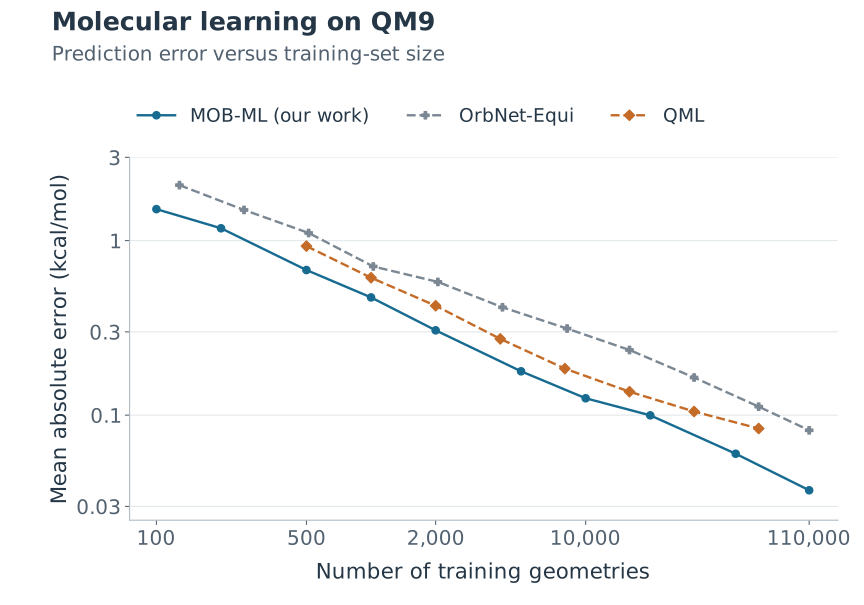
Molecular-orbital-based machine learning (MOB-ML) uses information from inexpensive electronic-structure calculations to describe the molecule. Molecular orbitals encode its electronic environment, providing a physically motivated basis for learning the correlation energy missing from a mean-field calculation. During my Ph.D. with Thomas Miller, I worked on the representations, physical constraints, and learning algorithms that make this approach accurate and transferable.
One important ingredient is size consistency: the energy of well-separated, noninteracting fragments should equal the sum of their individual energies. Incorporating this requirement improved transfer from small training molecules to much larger systems. Our studies examined diverse chemical settings, including organic molecules, transition-metal complexes, noncovalent interactions, and transition states, testing whether a common orbital-based representation could remain useful across different electronic environments.
I also extended MOB-ML to molecular dipole moments. Unlike an energy, a dipole is a vector: rotating the molecule should rotate the predicted dipole in the same way. We developed rotationally equivariant Gaussian-process regression with derivatives to incorporate this structure, allowing the framework to learn electronic-correlation contributions to dipoles while respecting their transformation under rotation.
Efficient training is equally important. Kernel addition Gaussian-process regression (KA-GPR) exploits the local decomposition of molecular energies while learning directly from total correlation energies. This improves data efficiency, supports open-shell and multireference applications, and expands practical training scale. Complementary work on scalable Gaussian-process algorithms and clustering of chemical space further reduces learning costs. Applications to potential energy surfaces connect these developments to molecular quantum dynamics, where many accurate energy evaluations are needed.
Selected publications
- Improved accuracy and transferability of molecular-orbital-based machine learning: Organics, transition-metal complexes, non-covalent interactions, and transition states. J. Chem. Phys. 154, 064108 (2021).
- Molecular dipole moment learning via rotationally equivariant Gaussian process regression with derivatives in molecular-orbital-based machine learning. J. Chem. Phys. 157, 104109 (2022).
Molecular-orbital-based machine learning for open-shell and multi-reference systems with kernel addition Gaussian process regression. J. Chem. Phys. 157, 154105 (2022).
- Accurate molecular-orbital-based machine learning energies via unsupervised clustering of chemical space. J. Chem. Theory Comput. 18, 4826–4835 (2022).
- Molecular energy learning using alternative blackbox matrix-matrix multiplication algorithm for exact Gaussian process. NeurIPS AI for Science Workshop (2021).
- Fast near ab initio potential energy surfaces using machine learning. J. Phys. Chem. A 126, 4013–4024 (2022).
Additional Research
Measurement-induced phase transitions
Quantum evolution can build entanglement, while measurements can suppress it; their competition produces measurement-induced phase transitions. With collaborators, I contributed to an experimental demonstration on superconducting quantum processors using scalable cross-entropy benchmarking. This work connects quantum many-body phenomena with protocols that can be implemented and tested on quantum hardware.
- Experimental demonstration of scalable cross-entropy benchmarking to detect measurement-induced phase transitions on a superconducting quantum processor. Phys. Rev. Lett. 134, 120401 (2025).
Molecular dynamics and materials
- Molecular quantum dynamics. I developed stable and efficient ring-polymer molecular-dynamics methods for capturing nuclear quantum effects. Related work applied these methods to bimolecular reaction rates. Integrators (2021) · Reaction rates (2022)
- Electron transport in semiconductors. I studied high-field transport and hot-electron noise in GaAs, using first-principles calculations and semi-analytical models to investigate two-phonon scattering. First-principles study (2022) · Semi-analytical model (2023)
- Electronic structure of graphene nanoribbons. I studied the regulation of electronic structure through long-range dopant–dopant coupling. Publication (2020)
See the complete publication list for all papers.